
nur für Forschungszwecke
Idarubicin HCl Topoisomerase Inhibitor
Kat.-Nr.: S1228
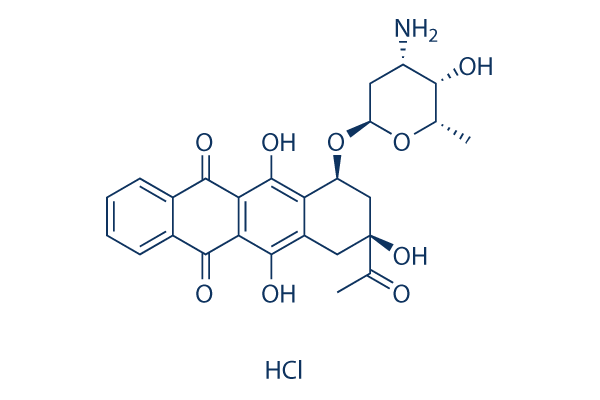
Chemische Struktur
Molekulargewicht: 533.95
Qualitätskontrolle (Quality Control)
| Verwandte Ziele | HDAC PARP ATM/ATR DNA-PK WRN DNA/RNA Synthesis PPAR Sirtuin Casein Kinase eIF |
|---|---|
| Weitere Topoisomerase Inhibitoren | Camptothecin (CPT) Betulinic acid (S)-10-Hydroxycamptothecin Beta-Lapachone Ellagic acid Amonafide Voreloxin (SNS-595) hydrochloride Hydroxy Camptothecine Cu(II)-Elesclomol 7-Ethylcamptothecin |
Zellkultur, Behandlung & Arbeitskonzentration
(Cell Culture, Treatment & Working Concentration)
| Zelllinien | Assay-Typ | Konzentration | Inkubationszeit | Formulierung | Aktivitätsbeschreibung | PMID |
|---|---|---|---|---|---|---|
| K562 erythroleukemic cells | Cytotoxic assay | Cytotoxic activity against K562 erythroleukemic cells, IC50=0.002 μM | 10425093 | |||
| K562 cells | Cytotoxic assay | 5 days | Cytotoxicity against human K562 cells after 5 days by XTT assay, IC50=0.012 μM | 18076140 | ||
| mouse DA-3 cells | Cytotoxic assay | 24 h | Cytotoxicity against mouse DA-3 cells after 24 hrs by XTT assay, IC50=2.3 μM | 24900668 | ||
| human ES2 cells | Cytotoxic assay | 24 h | Cytotoxicity against human ES2 cells after 24 hrs by XTT assay, IC50=3.2 μM | 24900668 | ||
| human K562 cells | Proliferation assay | 72 h | Antiproliferative activity against human K562 cells after 72 hrs, GI50=3.3 μM | 25420175 | ||
| human SKOV3 cells | Cytotoxic assay | 24 h | Cytotoxicity against human SKOV3 cells after 24 hrs by XTT assay, IC50=4.5 μM | 24900668 | ||
| HepG2 cells | Function assay | Inhibition of liver stage Plasmodium berghei infection in HepG2 cells, IC50=6.62 μM | 22586124 | |||
| human MCF7 cells | Cytotoxic assay | 24 h | Cytotoxicity against human MCF7 cells after 24 hrs by XTT assay, IC50=7.3 μM | 24900668 | ||
| mouse DA-3 cells | Cytotoxic assay | 20 μM | Cytotoxicity against mouse DA-3 cells assessed as reduction in cell viability at 20 uM by XTT assay | 24900668 | ||
| neural precursor cells | Function assay | Inhibition of neurosphere proliferation of mouse neural precursor cells by MTT assay | 17417631 | |||
| VERO-E6 | Function assay | 48 hrs | Toxicity CC50 against VERO-E6 cells determined at 48 hours by high content imaging (same conditions as 2_LEY without exposure to 0.01 MOI SARS CoV-2 virus), CC50=0.2μM | ChEMBL | ||
| Klicken Sie hier, um weitere experimentelle Daten zu Zelllinien anzuzeigen | ||||||
Chemische Informationen, Lagerung & Stabilität (Chemical Information, Storage & Stability)
| Molekulargewicht | 533.95 | Formel | C26H27NO9.HCl |
Lagerung (Ab dem Eingangsdatum) | |
|---|---|---|---|---|---|
| CAS-Nr. | 57852-57-0 | SDF herunterladen | Lagerung von Stammlösungen |
|
|
| Synonyme | 4-demethoxydaunorubicin (NSC256439, 4-DMDR) HCl | Smiles | CC1C(C(CC(O1)OC2CC(CC3=C2C(=C4C(=C3O)C(=O)C5=CC=CC=C5C4=O)O)(C(=O)C)O)N)O.Cl | ||
Löslichkeit (Solubility)
|
In vitro |
DMSO
: 100 mg/mL
(187.28 mM)
Water : 8 mg/mL Ethanol : Insoluble |
Molaritätsrechner
|
In vivo |
|||||
In-vivo-Formulierungsrechner (Klare Lösung)
Schritt 1: Geben Sie die untenstehenden Informationen ein (Empfohlen: Ein zusätzliches Tier zur Berücksichtigung von Verlusten während des Experiments)
Schritt 2: Geben Sie die In-vivo-Formulierung ein (Dies ist nur der Rechner, keine Formulierung. Bitte kontaktieren Sie uns zuerst, wenn es im Abschnitt "Löslichkeit" keine In-vivo-Formulierung gibt.)
Berechnungsergebnisse:
Arbeitskonzentration: mg/ml;
Methode zur Herstellung der DMSO-Stammlösung: mg Wirkstoff vorgelöst in μL DMSO ( Konzentration der Stammlösung mg/mL, Bitte kontaktieren Sie uns zuerst, wenn die Konzentration die DMSO-Löslichkeit der Wirkstoffcharge überschreitet. )
Methode zur Herstellung der In-vivo-Formulierung: Nehmen Sie μL DMSO Stammlösung, dann hinzufügenμL PEG300, mischen und klären, dann hinzufügenμL Tween 80, mischen und klären, dann hinzufügen μL ddH2O, mischen und klären.
Methode zur Herstellung der In-vivo-Formulierung: Nehmen Sie μL DMSO Stammlösung, dann hinzufügen μL Maisöl, mischen und klären.
Hinweis: 1. Bitte stellen Sie sicher, dass die Flüssigkeit klar ist, bevor Sie das nächste Lösungsmittel hinzufügen.
2. Achten Sie darauf, das/die Lösungsmittel der Reihe nach hinzuzufügen. Sie müssen sicherstellen, dass die bei der vorherigen Zugabe erhaltene Lösung eine klare Lösung ist, bevor Sie mit der Zugabe des nächsten Lösungsmittels fortfahren. Physikalische Methoden wie Vortex, Ultraschall oder ein heißes Wasserbad können zur Unterstützung des Lösens verwendet werden.
Wirkmechanismus (Mechanism of Action)
| Merkmale |
Idarubicin is a substrate for CYP450 2D6 and 2C9.
|
|---|---|
| Targets/IC50/Ki |
Topo II (MCF-7 cells)
(Cell-free assay) 3.3 ng/mL
Multicellular spheroids
(Cell-free assay) 7.9 ng/mL
|
| In vitro |
Idarubicin has significant cytotoxic activity against multicellular spheroids, comparable to the antiproliferative effects on monolayer cells. Idarubicin inhibits CYP450 2D6. Idarubicin is about 57.5-fold and 25-fold more active than doxorubicin and epirubicin, respectively. Idarubicin is able to overcome P-glycoprotein-mediated multidrug resistance. Idarubicin inhibits PMN superoxide radical formation. Idarubicin could be coupled to the monoclonal antibodies (anti-Ly-2.1, anti-L3T4, or anti-Thy-1) with retention of protein solubility and antibody activity. Idarubicin inhibits the proliferation of NALM-6 cells with an IC50 of 12 nM. |
| Kinase-Assay |
CYP450-Stoffwechsel-Experimente
|
|
Die Bewertung des Idarubicin-Metabolismus durch die CYP450-Isoenzyme 3A4, 2D6, 2C8, 2C9 und 1A2 wird unter Verwendung isolierter humaner CYP450-Proteine für jedes Isoform durchgeführt. Für diese Bewertungen wird die High-Throughput-P450-Inhibitions-Testmethode eingesetzt. Die Stoffwechsel-Experimente sind darauf ausgelegt, die folgenden Eigenschaften jedes Medikaments zu untersuchen: (1) ob Idarubicin ein Substrat der CYP450 3A4, 2C8, 2C9, 1A2 oder 2D6 Isoenzyme ist; (2) ob der Metabolismus durch bekannte Inhibitoren jedes Isoenzyms beeinflusst wird; (3) ob Idarubicin Inhibitoren von CYP450-Isoenzymen ist; und (4) ob Caspofungin oder Itraconazol den CYP450-Metabolismus von Idarubicin hemmen. Dibenzylfluorescein (DBF) (CYP3A4, CYP2C8, CYP2C9), 3-Cyano-7-ethoxycoumarin (Cyp1A2) und 7-Methoxy-4-(aminomethyl)-coumarin (MAMC) (CYP2D6) sind die bekannten Substrate, die als Kontrollen verwendet werden, um die jeweilige Isoenzymaktivität zu bestätigen und die Auswirkungen von Idarubicin auf die Isoenzymaktivität zu bewerten. Darüber hinaus werden Ketoconazol, Quercetin, Suflaphenazol, Furafyllin und Chinidin als Kontroll-CYP450-Inhibitoren für die 3A4, 2C8, 2C9, 1A2 bzw. 2D6 Isoenzyme verwendet. Das Substrat, der Inhibitor plus Idarubicin wie angegeben werden zu jeder Proteinprobe hinzugefügt und für 20 bis 60 Minuten, wie vom Hersteller empfohlen, bei 37 °C inkubiert. Die Reaktionen werden mit einer organischen Lösung gestoppt, und die Proben werden dann entsprechend mittels Fluoreszenz-Plattenlesegerät analysiert. Für jedes Experiment werden Kontrollproben mit einer bekannten Menge Substrat und synthetisiertem Metaboliten, in Abwesenheit des Isoenzyms, für qualitative Vergleiche hergestellt. Alle Experimente werden in dreifacher Ausführung durchgeführt.
|
|
| In vivo |
Reduction of Idarubicin is dependent upon ketone reductases, and proceeds more stereoselectively than that of most ketones giving rise to the (13S)-epimer almost exclusively. The high stereospecificity in Idarubicin reduction might result from chiral induction due to the presence of asymmetric centres near to the carbonyl group in Idarubicin. |
Literatur |
|
Klinische Studieninformationen (Clinical Trial Information)
(Daten von https://clinicaltrials.gov, aktualisiert am 2024-05-22)
| NCT-Nummer | Rekrutierung | Erkrankungen | Sponsor/Kooperationspartner | Startdatum | Phasen |
|---|---|---|---|---|---|
| NCT05697510 | Recruiting | Acute Myeloid Leukemia (AML) |
Nantes University Hospital |
March 16 2023 | Phase 1 |
| NCT02652871 | Completed | Leukemia |
M.D. Anderson Cancer Center|Eli Lilly and Company|High Impact Clinical Research Support Program |
May 9 2016 | Phase 1 |

Produkte sind nur für Forschungszwecke bestimmt. Nicht für den menschlichen Gebrauch. Wir verkaufen nicht an Patienten.
©Copyright 2013 Selleck Chemicals. Alle Rechte vorbehalten.